Flag ranges that start a quintuplet in which two overlapping inversions moved the three central ranges, and the two flanking ranges did not move.
Arguments
- gb
A
GBreaks()object.- details
Returns more metadata columns if
TRUE.
Details
With the five target ranges named X A B C Y in their order on the
target genome, and lower case meaning the minus strand, the query
genome reads one of the following:
X b c A Y    ABC -> baC -> bcA
X C a b Y    ABC -> Acb -> Cab
y B A c x    cba -> BCa -> BAc
y a C B x    cba -> cAB -> aCB
The last two are the first two read on the other strand of the query
genome. The flanks X and Y must be on the strand shown: with flanks on
the opposite strand, the same central triplet requires three inversions.
The flag is set on range X, as flagInversions() sets its flag on the
left flank of an inversion. The five ranges must be on the same target
sequence, and their strand must be known (not *).
See also
Other Flagging functions:
flagAll(),
flagColinearAlignments(),
flagInversions(),
flagLongShort(),
flagPairs(),
flagTranslocations(),
flagTwinInversions()
Other Inversion functions:
filterDoubleInversions(),
filterInversions(),
flagInversions(),
flagPairs(),
flagTwinInversions(),
flipInversions(),
leftInversionGaps(),
removeDoubleInversions(),
removeInversions(),
showInversions()
Other Structural variants:
StructuralVariants,
flagInversions(),
flagPairs(),
flagTranslocations(),
flagTwinInversions(),
plotApairOfChrs()
Examples
# Start colinear. Lower case meands minus strand
z0 <- GBreaks(target = GRanges(c(A="T:10-15:+", B="T:20-25:+", C="T:30-35:+")),
query = GRanges(c(A="Q:10-15", B="Q:20-25", C="Q:30-35")))
# Swap coordinates of A and B on the query and flip strands ABC -> baC
z1 <- GBreaks(target = GRanges(c(A="T:10-15:-", B="T:20-25:-", C="T:30-35:+")),
query = GRanges(c(a="Q:20-25", b="Q:10-15", C="Q:30-35")))
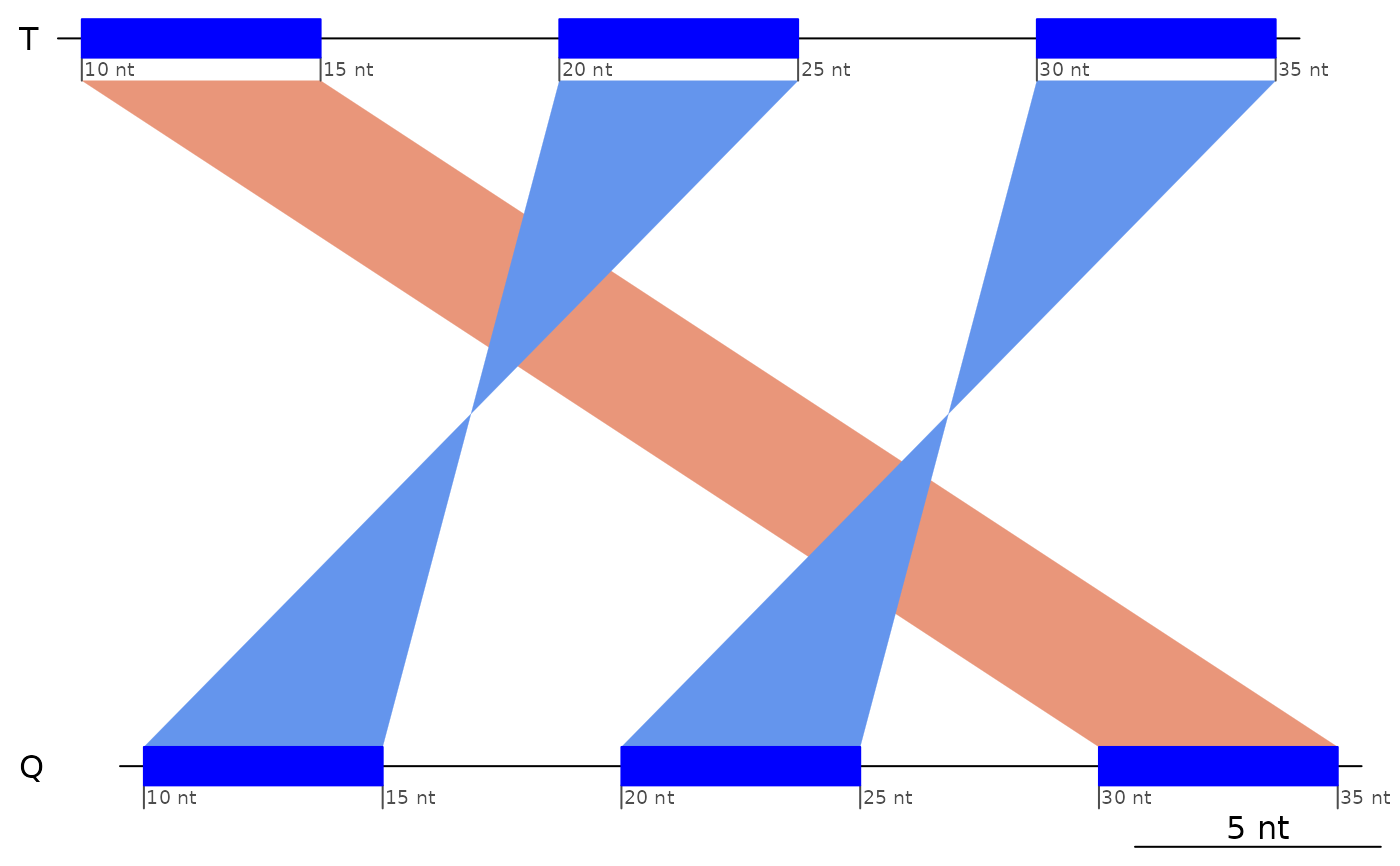
# Now query order is b - a - C. Swap a and C and flip strands baC -> bcA
z2 <- GBreaks(target = GRanges(c(A="T:10-15:+", B="T:20-25:-", C="T:30-35:-")),
query = GRanges(c(A="Q:30-35", b="Q:10-15", c="Q:20-25")))
# Altogether, there are:
# ABC -> baC -> bcA
# ABC -> Acb -> Cab
# cba -> BCa -> BAc
# cba -> cAB -> aCB
# z2 has same topoloty as package example
plotApairOfChrs(z2)
 plotApairOfChrs(exampleDoubleInversion1)
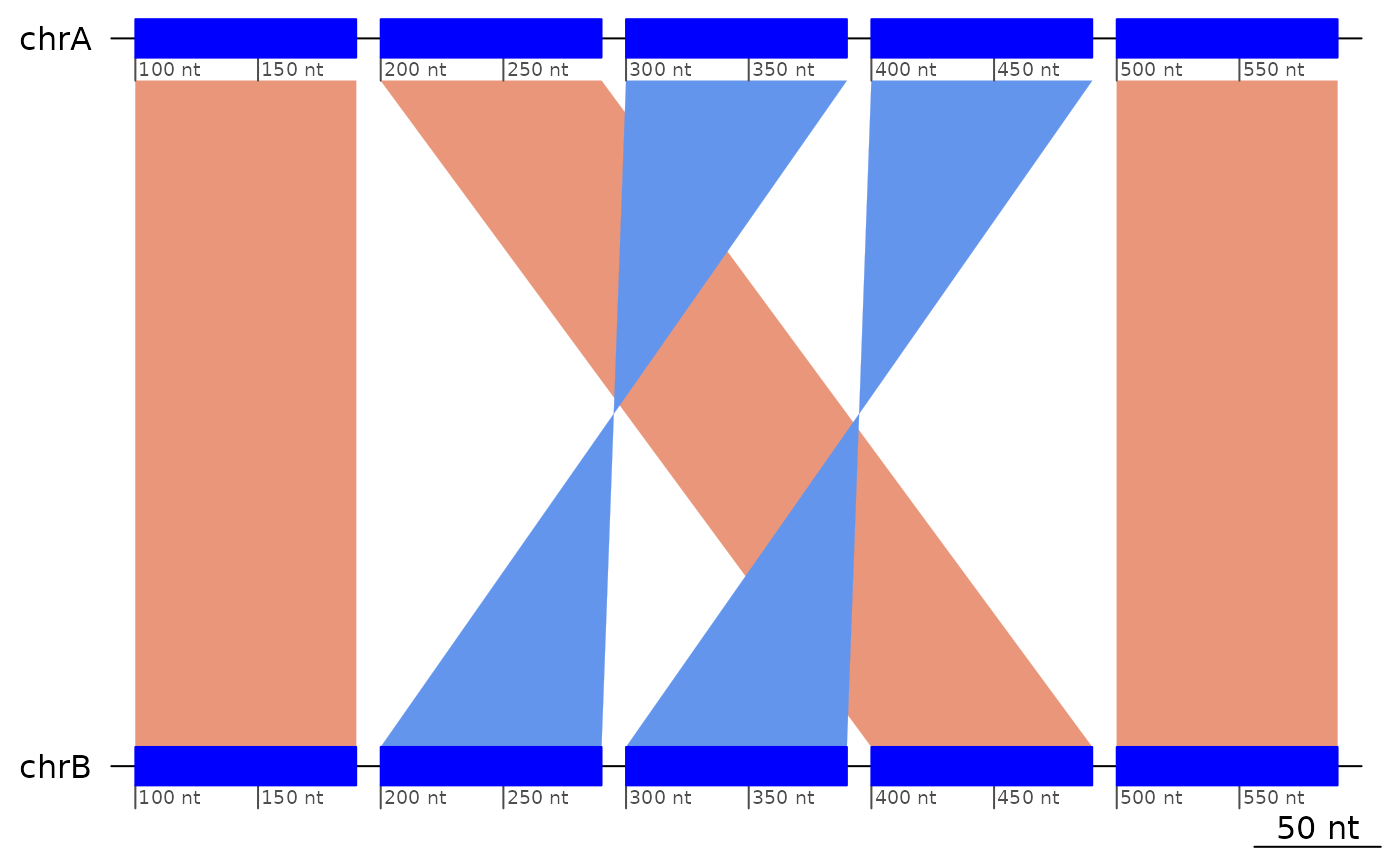 flagDoubleInversions(exampleDoubleInversion1 )
#> GBreaks object with 5 ranges and 2 metadata columns:
#> seqnames ranges strand | query Dbl
#> <Rle> <IRanges> <Rle> | <GRanges> <logical>
#> [1] chrA 100-190 + | chrB:100-190 TRUE
#> [2] chrA 200-290 + | chrB:400-490 FALSE
#> [3] chrA 300-390 - | chrB:200-290 FALSE
#> [4] chrA 400-490 - | chrB:300-390 FALSE
#> [5] chrA 500-590 + | chrB:500-590 FALSE
#> -------
#> seqinfo: 1 sequence from an unspecified genome
exampleDoubleInversion1Rev <- reverse(exampleDoubleInversion1) |> sort(ignore.strand = TRUE)
exampleDoubleInversion1Rev[2:4] |> plotApairOfChrs()